Key Outcomes:
- Survodutide (Boehringer Ingelheim) – P3 SYNCHRONIZE-2, obesity and T2D: Met both co-primary endpoints at Wk 76; achieved up to 13.1% weight loss and 1.21 percentage-point HbA1c reduction vs pbo. GI-related discontinuations were higher (18% vs 1.2%)
- Rina-S (Genmab) – P1/2 RAINFOL-01, platinum-resistant ovarian cancer: Demonstrated a 45.9% cORR, 12.1 mos mDoR and 9.5 mos mPFS; four P3 studies are planned or underway across ovarian and endometrial cancer settings
- Vamikibart (Roche) – P3 MEERKAT/SANDCAT, uveitic macular edema: 1-yr data showed sustained vision improvements and reduced macular thickness; the US FDA accepted the BLA, with a decision expected by Jul 2027
- Efzimfotase alfa (AstraZeneca/Alexion) – P3 HICKORY, hypophosphatasia: Missed the primary 6-minute walk test endpoint at Wk 25 (p=0.1675), despite numerical improvement vs pbo; fatigue improved nominally significantly, and the US FDA granted Priority Review
- Admilparant (Bristol Myers Squibb) – P3 ALOFT-IPF, idiopathic pulmonary fibrosis: The 60 mg twice-daily dose met the primary endpoint for FVC change at Wk 52; the 120 mg dose did not reach statistical significance. P3 ALOFT-PPF results are expected in early 2027
- ABO2203 (Abogen/Novartis) – mRNA-encoded T-cell engager, autoimmune diseases: Novartis secured an exclusive worldwide license; Abogen is eligible for $575M upfront and up to approx. $7.2B in milestone payments, plus potential royalties
- Ivonescimab (AstraZeneca/Summit Therapeutics) – solid tumors: AstraZeneca completed a $2B strategic equity investment to accelerate development across solid tumors, including combinations with ADCs; the US FDA’s target action date for the EGFR-mutated advanced NSCLC application is Nov 14, 2026
- LIBERVANT/diazepam buccal film (Eton Pharmaceuticals/Aquestive) – seizure clusters: Eton secured exclusive US commercialization rights; launch is anticipated in Apr 2027, subject to final US FDA approval. The supplemental application for ages 6–11 has a Mar 29, 2027 target action date
- Infigratinib (BridgeBio) – achondroplasia in children: Received US FDA Priority Review, with a Feb 4, 2027 target action date; P3 PROPEL 3 met its primary endpoint, showing a +2.10 cm/year treatment difference in annualized height velocity vs pbo
- Rhapsido/remibrutinib (Novartis) – symptomatic dermographism: Received US FDA approval as the first specifically approved treatment for adults inadequately controlled by H1 antihistamines; complete hive response at Wk 12 was 29.3% vs 14.0% with pbo in the P3 RemIND study
- KERENDIA/finerenone (Bayer) – CKD without diabetes: US FDA accepted the sNDA based on P3 FIND-CKD; treatment slowed annual eGFR decline (-3.3 vs -4.0 mL/min/1.73 m²/year) and reduced the risk of the composite kidney-cardiovascular outcome by 23% vs pbo
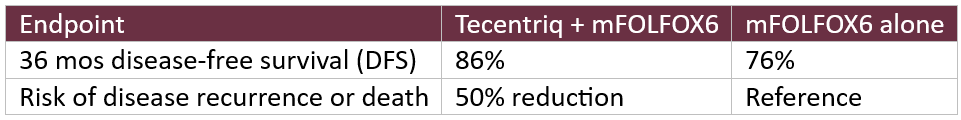
- Tecentriq/atezolizumab (Roche) – resected stage III dMMR colon cancer: Received US FDA approval for adjuvant uses with chemotherapy; P3 ATOMIC showed 86% DFS at 3 yrs vs 76% with chemotherapy alone, with a 50% reduction in recurrence or death risk
- Souffle Therapeutics – Series B financing: Raised $135M to advance three lead programs toward clinical studies, targeting facioscapulohumeral muscular dystrophy, heart failure with reduced ejection fraction and an undisclosed muscle/metabolic disorder
- OMass Therapeutics – Series C financing: Raised $310M to advance OMS1620 through P2 for congenital adrenal hyperplasia and ACTH-dependent Cushing’s syndrome, and generate initial clinical data for an SLC15A4 inhibitor targeting lupus and related diseases
October 01, 2026
Boehringer Ingelheim’s Survodutide Meets Co-primary Endpoints in P3 SYNCHRONIZE-2 Trial in Obesity and T2D
BI’s investigational dual glucagon/GLP-1 receptor agonist Survodutide (BI 456906) met both co-primary endpoints in wk 76 P3 SYNCHRONIZE-2 study involving 755 adults with obesity or overweight and type 2 diabetes (T2D). Results were presented at EASD 2026 and published in The NEJM. BI is continuing to evaluate Survodutide across obesity, MASH and T2D, with SYNCHRONIZE-CVOT study results expected later in 2026, while the P3 SYNCHRONIZE-T2Dstudy will further evaluate glycemic control in adults with T2D.
Efficacy results at Wk 76:
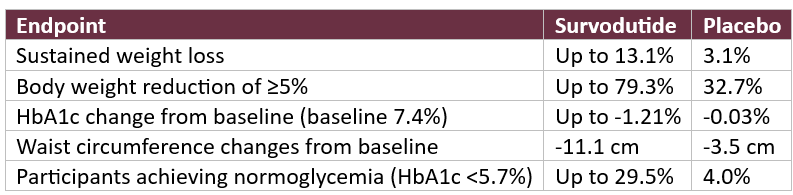
All listed comparisons were statistically significant (p<0.0001), as reported in the source.
Safety Profile: Gastrointestinal adverse events, including nausea, vomiting, diarrhea, and constipation, were the most common and were mostly mild to moderate. Discontinuation due to gastrointestinal adverse events occurred in 18% of Survodutide-treated participants vs 1.2% with pbo.
Source: GlobeNewswire PR Oct 1, 2026
October 03, 2026
Genmab’s Rina-S Demonstrates Positive ORR in P1/2 RAINFOL-01 Study in Platinum-Resistant Ovarian Cancer
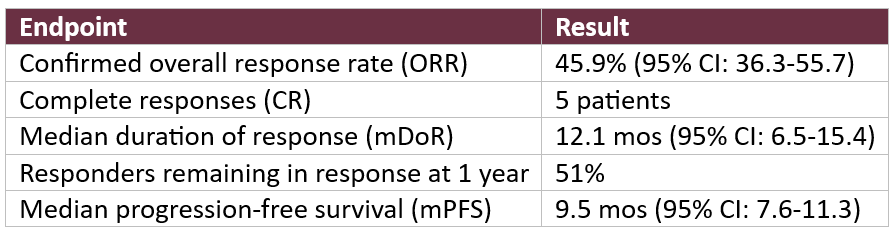
Genmab reported results from Part C of the P1/2 RAINFOL™-01 study evaluating Rinatabart sesutecan (Rina-S; GEN1184), an investigational FRα-targeted, TOPO1-inhibitor ADC, as monotherapy in platinum-resistant ovarian cancer (PROC). Among 109 treated patients receiving 120 mg/m² Q3W, Rina-S demonstrated a confirmed ORR of 45.9%, including 5 CRs, a mDoR of 12.1 mos and mPFS of 9.5 mos. Antitumor activity was observed regardless of FRα expression, including in patients with low or no expression, and regardless of prior mirvetuximab treatment. Results were presented at the 2026 International Gynecologic Cancer Society (IGCS) Congress in Montreal, Canada. Genmab is advancing Rina-S in 4 P3 studies across PROC (RAINFOL-02), endometrial cancer (RAINFOL-03), PSOC maintenance therapy (RAINFOL-04)and 2L PSOC (RAINFOL-07). It is also being evaluated in the P1/2 RAINFOL-01 study and P2 RAINFOL-05 study in NSCLC and RAINFOL-09study in advanced gastrointestinal cancers.
Efficacy results:
Safety Profile: The most common TEAEs included fatigue, nausea, vomiting, constipation, decreased appetite, and abdominal pain, mostly Gr 1-2 gastrointestinal events. Hematologic TEAEs included anaemia, neutropenia, decreased platelet count and thrombocytopenia. SAEs occurred in approximately one-third of participants, while 5.5% discontinued treatment due to TEAEs. No ocular toxicity, peripheral neuropathy, interstitial lung disease, or stomatitis safety signals were observed.
Source: GlobeNewswire PR Oct 3, 2026
October 10, 2026
Roche Reports Sustained Vision Improvements with Vamikibart in P3 Uveitic Macular Edema Studies
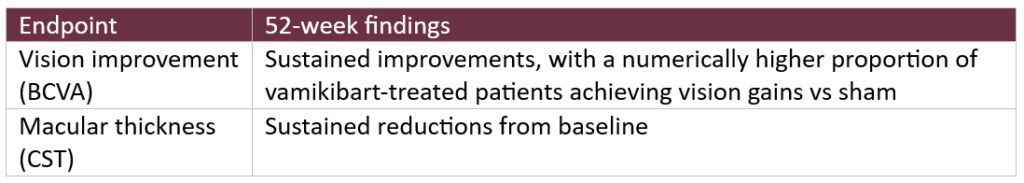
Roche presented 1-yr data from the P3 MEERKAT and SANDCAT studies evaluating intravitreal Vamikibart, an investigational anti IL-6 mAb, vs sham injections in adults with uveitic macular edema (UME). Across both studies, Vamikibart demonstrated a numerically higher proportion of vamikibart-treated patients achieved vision gains vs sham at wk. 52, with sustained improvements in best corrected visual acuity (BCVA) and reductions in central subfield thickness (CST) from baseline. Approx. two-thirds of eligible patients required no retreatment after wk. 16, while most patients requiring retreatment received only one additional injection. The data were presented at the AAO’26 in New Orleans, LA. The US FDA accepted Roche’s BLA, with a decision expected by Jul’27; regulatory submissions have also been filed and accepted in the EU, China and Japan.
Efficacy results
Safety Profile: Vamikibart was well tolerated, with a low incidence of treatment-related ocular adverse events and intraocular inflammation.
Source: GlobeNewswire PR Oct 10, 2026
October 11, 2026
AstraZeneca Reports P3 HICKORY Results for Efzimfotase Alfa in Hypophosphatasia
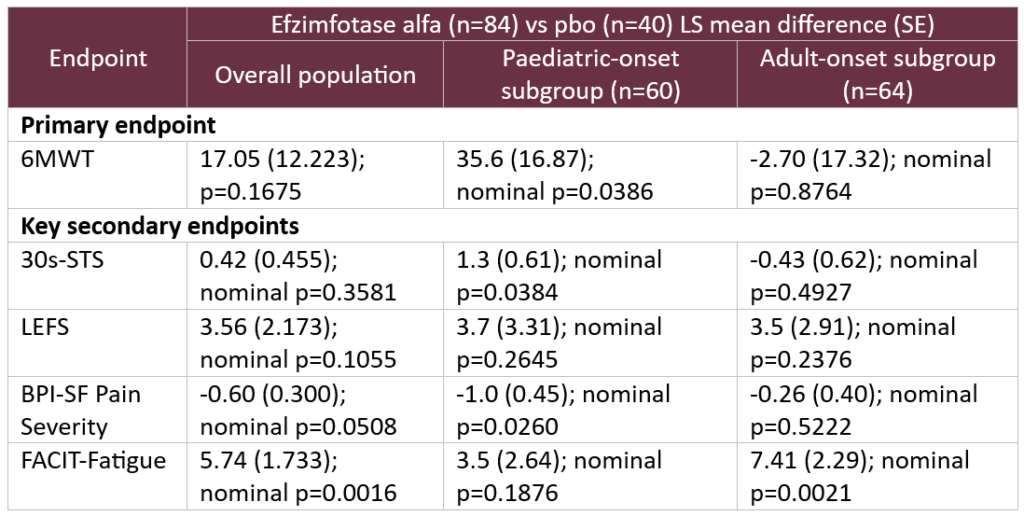
AZ’s Alexion reported results from P3 HICKORY study evaluating efzimfotase alfa (ALXN1850), an investigational enzyme replacement therapy, in treatment naive adolescents and adults aged ≥12 years with hypophosphatasia (HPP) who had not previously received Strensiq (asfotase alfa). At wk. 25, efzimfotase alfa didn’t achieve statistical significance on the primary endpoint of 6MWT, despite a numerical improvement of 17.05 metres vs pbo (p=0.1675). The therapy demonstrated a nominally significant improvement in the key secondary endpoint of FACIT-Fatigue, with an LS mean difference of 5.74 points vs pbo (nominal p=0.0016), while other secondary endpoints numerically favoured treatment. In a post hoc analysis, 45.2% of treated patients achieved a minimum clinically important difference (MCID) in 6MWT vs 30% with pbo (nominal p=0.0363). In the paediatric onset subgroup, nominally significant improvements were observed in 6MWT, 30-second Sit-to-Stand (30s-STS) and BPI-SF Pain Severity, while the adult-onset subgroup showed a nominally significant improvement in FACIT-Fatigue. Improvements across mobility, physical function, pain and fatigue continued through wk. 49 OLE. HICKORY results were presented at the 2026 American Society for Bone and Mineral Research (ASBMR) annual meeting in Boston. The US FDA recently granted Priority Review to efzimfotase alfa for HPP.
Efficacy results at Wk 25:
Safety Profile: Efzimfotase alfa was generally well tolerated. In a pooled analysis of the MULBERRY and HICKORY studies, the annualised injection-site reaction rate was 5 times lower than that reported with Strensiq in its registrational studies during the first 24 wks. Patients receiving efzimfotase alfa had a median of 98.8% injection-site reaction-free days.
Source: AstraZeneca PR Oct 11, 2026
October 12, 2026
BMS’s Admilparant Meets Primary Endpoint in P3 IPF Study
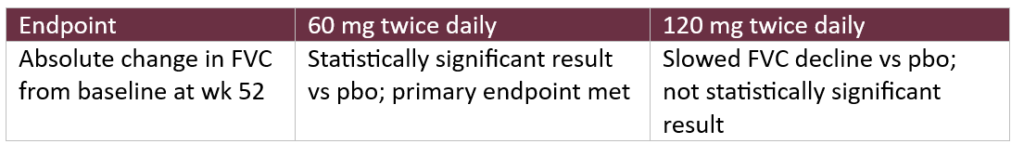
Bristol Myers Squibb (BMS) reported positive topline results from the P3 ALOFT-IPF study, evaluating the efficacy, safety and tolerability of oral admilparant (BMS-986278), a potential FIC lysophosphatidic acid receptor 1 (LPA1) antagonist in adults with idiopathic pulmonary fibrosis (IPF). The trial evaluated two dose levels of admilparant, 60 mg & 120 mg, BID for ≥52 weeks vs pbo. The study met its primary endpoint at 60 mg dose, demonstrating a statistically significant improvement vs pbo in absolute change in forced vital capacity (FVC) from baseline at wk 52. The 120 mg dose slowed FVC decline but did not achieve statistical significance. BMS will evaluate P3 ALOFT-IPF data and engage with health authorities regarding regulatory submissions, with detailed results expected at a future medical meeting. Admilparant topline results from the P3 ALOFT-PPF study in progressive pulmonary fibrosis (PPF) are also expected in early 2027.
Efficacy results:
Safety Profile: Admilparant demonstrated a favourable safety and tolerability profile at both doses. Discontinuations due to AEs were low and similar to pbo.
October 06, 2026
BridgeBio Receives US FDA Priority Review for Oral Infigratinib NDA in Children with Achondroplasia
BridgeBio Pharma’s NDA for investigational oral infigratinib, an FGFR3 inhibitor for children with achondroplasia, received Priority Review by the US FDA, with a PDUFA target action date of Feb 4, 2027. The regulatory submission is supported by the P3 PROPEL 3 study, which met its primary and key secondary endpoints at wk 52, demonstrating a mean annualized height velocity (mAHV) treatment difference of +2.10 cm/year (p<0.0001) and a significant improvement in height Z-score. Exploratory findings included statistically significant improvements in body proportionality among children <8 yrs and in arm span Z-score. Favourable trends were also observed in sleep apnea and otitis media events. Treatment was well tolerated, with no study-drug-related SAEs or discontinuations. Results were published in The NEJM and presented at the 2026 International Congress of Children’s Bone Health (ICCBH). Additional data presented at ESPE explored potential benefits beyond height, including stabilizing sleep apnea measures, reducing rate of ear infections and impacting body composition for people with achondroplasia. BridgeBio anticipates a US launch upon approval and plans to submit a MAA for achondroplasia to the EMA in Q4 2026. Infigratinib has also received BTD (based on PROPEL 2 study), ODD, FTD and RPDD from the US FDA, and ODD from the EMA.
Source: GlobeNewswire PR Oct 6, 2026
October 07, 2026
Novartis Receives US FDA Approval for Rhapsido (Remibrutinib) in Symptomatic Dermographism
Novartis received US FDA approval for Rhapsido® (remibrutinib), an oral, highly selective BTK inh., for adults with symptomatic dermographism (SD) inadequately controlled by H1 antihistamines, making it the first treatment approved by the US FDA specifically for SD. The approval was supported by the P3 RemIND study, in which significantly more patients achieved complete hive response at wk 12 with remibrutinib vs pbo (29.3% vs 14.0%; p=0.0229), with responses observed as early as wk 2. In clinical trials up to wk 24, Rhapsido demonstrated a consistent safety profile in SD & CSU; common AEs included nasal congestion, sore throat, nasopharyngitis, bleeding, headache, nausea and abdominal pain. The approval makes Rhapsido the only treatment approved in the US for both CSU and SD. Novartis plans to submit the full RemIND dataset to global health authorities where appropriate, while continuing to evaluate remibrutinib in other immune-mediated conditions, including hidradenitis suppurativa, food allergy and relapsing multiple sclerosis.
Source: Novartis PR Oct 7, 2026
October 08, 2026
US FDA Accepts Bayer’s sNDA for KERENDIA® in CKD Without Diabetes
Bayer’s sNDA for KERENDIA® (finerenone), a non-steroidal MRA, has been accepted by the US FDA for the treatment of adults with CKD without diabetes who are receiving background SoC. The application is based on the P3 FIND-CKD study, involving 1,584 adults, which met its primary endpoint by demonstrating a slower annual eGFR decline with finerenone vs pbo through month 32. KERENDIA also significantly reduced the risk of a composite kidney-cardiovascular outcome. If approved, the new indication would expand KERENDIA’s use to patients with CKD without diabetes, complementing its existing indications in CKD associated with T1D & T2D and heart failure with LVEF ≥40%.
Efficacy Results:
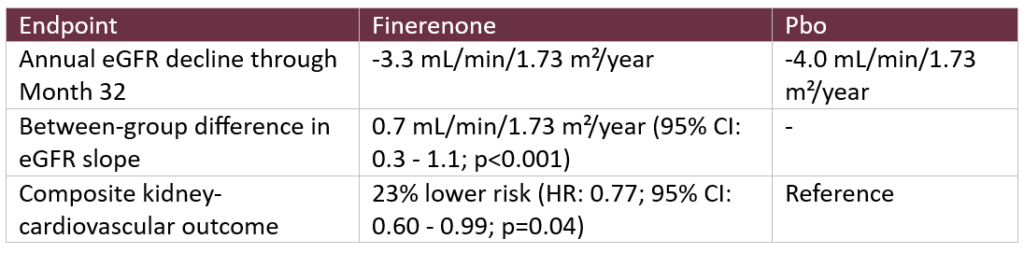
The composite outcome included kidney failure, sustained eGFR decrease ≥57%, hospitalization for heart failure or cardiovascular death.
Safety Profile: SAEs occurred at similar rates with finerenone and pbo (20.9% vs 21.2%). Hyperkalemia was more frequent with finerenone (17% vs 13.3%); serious hyperkalemia events occurred in less than 1% of patients.
Source: Business Wire PR Oct 8, 2026
October 09, 2026
Roche Receives US FDA Approval for Tecentriq-Based Adjuvant Regimen in Stage III dMMR Colon Cancer
Roche received US FDA approval for Tecentriq (atezolizumab) and Tecentriq Hybreza (atezolizumab and hyaluronidase-tqjs), in combination with a fluoropyrimidine and oxaliplatin, for the adjuvant treatment of stage III dMMR colon cancer following surgery. The approval was supported by the P3 ATOMIC study, which demonstrated combination of atezolizumab & mFOLFOX6 reduced the risk of disease recurrence or death by 50% vs CT alone. The regimen achieved 86% DFS at 36 mos vs 76% with CT alone, marking the first approved adjuvant immunotherapy regimen for stage III dMMR colon cancer and Tecentriq’s 12th indication in the US. Roche is pursuing additional regulatory submissions, incl. in EU.
Efficacy results:
Safety Profile: The safety profile of Tecentriq combined with mFOLFOX6 was consistent with previous studies.
Source: Roche PR Oct 09, 2026
October 02, 2026
Abogen Licenses mRNA-Encoded T-Cell Engager ABO2203 to Novartis in Up to $7.2B Deal
Abogen Biosciences entered into a licensing and option agreement with Novartis to advance RNA-encoded therapeutics, including an exclusive worldwide license to ABO2203, an investigational mRNA-encoded CD19xCD3 T-cell engager for B-cell mediated autoimmune diseases. ABO2203 is designed to reset B cells by utilizing mRNA to direct the endogenous production of T-cell engagers in vivo. The agreement also grants Novartis exclusive options to license a number of next-generation therapeutic assets developed on Abogen’s RNA platform. Abogen will receive $575M upfront and is eligible for up to approx.. $7.2B in milestone payments if all program options are exercised and agreed development, regulatory and commercial milestones are achieved, plus potential royalties on future product sales. The transaction remains subject to customary closing conditions, including regulatory clearances.
Source: PR Newswire PR Oct 2, 2026
October 05, 2026
AstraZeneca Completes $2B Strategic Equity Investment in Summit Therapeutics to Advance Ivonescimab
AstraZeneca completed its $2B strategic equity investment in Summit Therapeutics to accelerate development of ivonescimab (SMT112), an investigational PD-1/VEGF BsAb, across multiple solid tumors. The investment involved approx. 108,955 preferred shares, convertible into common shares at a 1:1,000 ratio, implying a common share price of $18.36/share, representing a premium to Summit’s closing share price on the announcement date. The funds will support accelerated development of ivonescimab across solid tumors, incl. combination studies with ADCs such as sonesitatug vedotin (Sone-Ve) and datopotamab deruxtecan (Datroway). Ivonescimab is being evaluated in indications including NSCLC, CRC and urothelial carcinoma. The BLA for ivonescimab in EGFR-mutated advanced NSCLC, supported by the P3 HARMONi study, was accepted for filing by the US FDA in Jan 2026, with a PDUFA target date of Nov 14, 2026. In 2026, Summit also planned to advance ivonescimab in urothelial carcinoma through the P2/3 HARMONi-GU1 study, with global clinical site activations planned to begin in Q4 2026. Ivonescimab is approved in China but remains investigational in Summit’s licensed territories, including the US and Europe.
Source: Business Wire PR Oct 5, 2026
October 14, 2026
Eton Pharmaceuticals Secures Exclusive US Licensing Rights to Aquestive’s LIBERVANT for Seizure Clusters
Eton Pharmaceuticals entered into an exclusive US licensing and supply agreement with Aquestive Therapeutics for LIBERVANT® (diazepam) buccal film, an orphan designated treatment for seizure clusters in patients with epilepsy. LIBERVANT has received tentative US FDA approval for patients aged 2-5 years & ≥12 years, while a supplemental application to expand the indication to patients aged 6-11 years is under review, with a PDUFA target action date of Mar 29, 2027. Full approval for patients aged ≥12 years is expected to become possible following the expiration of VALTOCO’s ODD exclusivity on Jan 10, 2027. Eton anticipates commercial launch in Apr’27, subject to final US FDA approval. The agreement expands Eton’s portfolio into neurology and provides exclusive US commercialization rights to a product targeting a seizure rescue market valued at > $450M annually. LIBERVANT has issued patent protection extending into 2038, with additional patent applications pending with the USPTO.
Source: GlobeNewswire PR Oct 14, 2026

October 13, 2026
Souffle Therapeutics Raises $135M in Series B Financing to Advance Three Lead Programs Toward Clinical Development
Souffle Therapeutics closed an oversubscribed $135M Series B financing led by Juvia Group, with participation from new investors TCG Crossover Management, BB Biotech, Fidelity Management & Research Company, Balyasny Asset Management and a sovereign wealth fund, alongside existing investors including Amgen Ventures, ARCH Venture Partners, Bessemer Venture Partners, Breyer Capital, The Invus Group, Janus Henderson, Leaps by Bayer, Polaris Partners, Safar Partners, TAS Partners, Temasek and Vida Ventures. The proceeds will support the advancement of three lead programs toward clinical studies and expansion of its pipeline across additional cell types and tissues. The lead program targets striated muscle for facioscapulohumeral muscular dystrophy (FSHD), while the second targets the phospholamban (PLN) gene in cardiomyocytes for heart failure with reduced ejection fraction (HFrEF); the third program targets an undisclosed area of muscle and metabolic disorders. The company’s REMEDY™ platform identifies tissue-specific internalizing receptors and develops proprietary binders to enable targeted therapeutic delivery.
Source: Souffle Therapeutics PR Oct 13, 2026
October 15, 2026
OMass Therapeutics Raises $310M in Series C Financing to Advance Endocrinology and Immunology Pipeline
OMass Therapeutics raised an oversubscribed $310M (£230M) Series C financing co-led by new investors Bain Capital Life Sciences, Forbion and Novo Holdings, with participation from additional new and existing investors. The proceeds will support the advancement of OMS1620, a melanocortin-2 (MC2) receptor antagonist, through P2 development for congenital adrenal hyperplasia (CAH) and ACTH-dependent Cushing’s syndrome. The financing will also support the generation of initial clinical data for an SLC15A4 inhibitor being developed for lupus and other diseases associated with interferon overproduction and pathogenic B-cell activation.
Source: GlobeNewswire PR Oct 15, 2026
Thank you for your attention! For any queries, contact us at bd@octavusconsulting.com
Interested in the presentation? Download Here
Babli Singh is a Senior Analyst with strong analytical skills and a deep understanding of the biopharmaceutical and healthcare industries. She specializes in monitoring emerging trends and distilling complex information into concise, insightful summaries that deliver clear value and actionable intelligence for a global professional audience.